
Материал
Medical CE & FDA Technical Compliance Register: Global Directives Update Monitoring Schedule
CLINICAL SAFETY POSITIONING
The Global Directives Update Monitoring Schedule constitutes a closed-loop regulatory intelligence architecture engineered to sustain continuous conformity across all active medical aesthetic device platforms. This register formalizes the surveillance cadence, verification pathways, and documentation controls required to maintain alignment with evolving international directives, harmonized standards, and national registration frameworks. The schedule is deployed as a governed compliance asset, not a static reference, and is auditable at every revision interval.
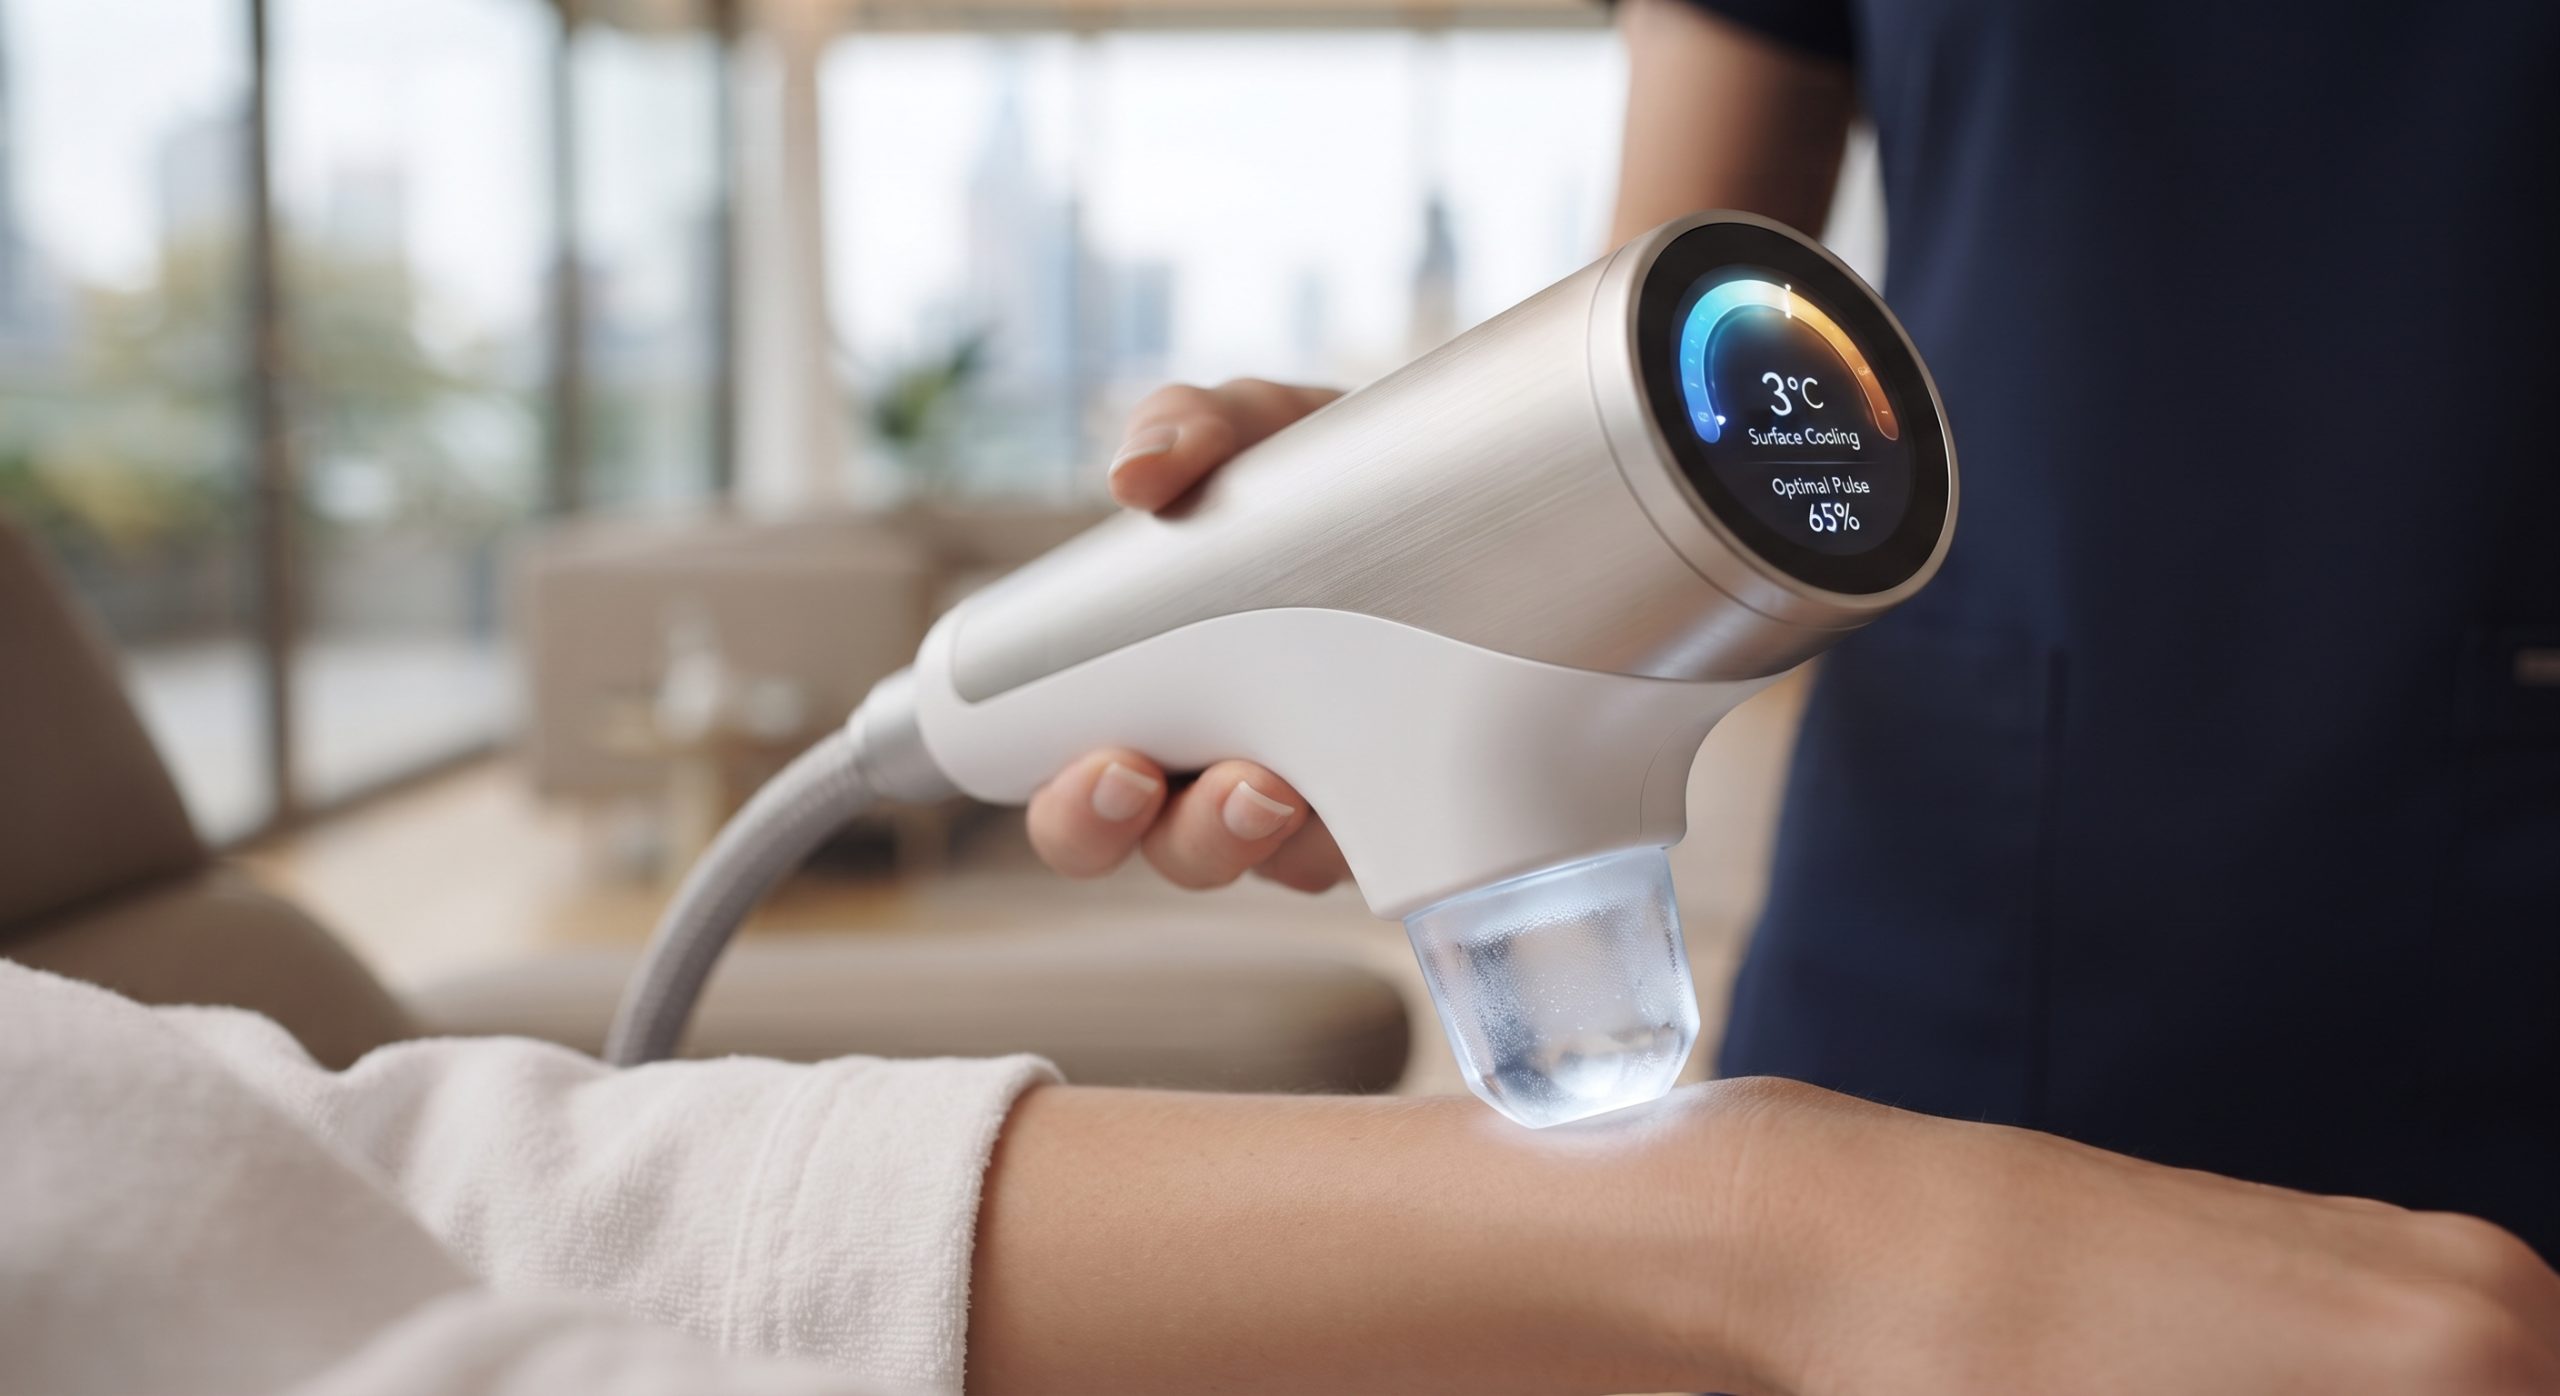
TEMPERATURE MONITORING CAPABILITIES
Directive convergence is monitored across four synchronized intelligence channels: (1) Notified Body communications and NANDO database status changes; (2) FDA CDRH database updates, including 510(k) and De Novo precedents; (3) ISO and IEC harmonized standard revisions affecting IEC 60601-1, IEC 60601-1-2, and ISO 13485; and (4) regional competent authority bulletins for EU MDR, UKCA, Health Canada, TGA, and ANVISA jurisdictions. Each channel is assigned a surveillance frequency tier and a named compliance owner.
PREMIUM COMPONENT ASSEMBLY
The monitoring schedule is executed through a structured compliance stack comprising a regulatory intelligence subscription layer, a change-impact assessment protocol, a documentation control system, and a corrective action gateway. Every detected directive change is logged with a unique reference identifier, impact classification (Critical / Major / Minor), affected technical file sections, and a mandatory response window. Traceability is preserved from source publication through to technical file amendment and, where required, notified body consultation.
DETAILED PARAMETERS
Surveillance intervals are tiered by directive volatility and jurisdictional risk exposure. The schedule defines mandatory review cycles, escalation triggers, and document retention periods consistent with ISO 13485 clause 4.2.5 and EU MDR Article 10(9) obligations. Post-Market Surveillance and Post-Market Clinical Follow-up inputs are formally linked to the directive monitoring loop to ensure that field data informs regulatory re-assessment.
| Compliance Parameter | Specification |
|---|---|
| Primary Regulatory Framework | EU MDR 2017/745 / FDA 21 CFR Part 820 |
| Quality Management Standard | ISO 13485:2016 |
| Risk Management Standard | ISO 14971:2019 |
| Electrical Safety Standard | IEC 60601-1:2005+A1+A2 |
| EMC Standard | IEC 60601-1-2:2014+A1 |
| Usability Engineering | IEC 62366-1 |
| Software Lifecycle | IEC 62304 |
| Surveillance Review Cadence | Quarterly (Critical) / Semi-Annual (Major) / Annual (Minor) |
| Escalation Response Window | 5 business days from classified change |
| Document Retention Period | 10 years minimum or device lifetime |
| Post-Market Surveillance Linkage | PMS / PMCF integrated into directive loop |
| Master Register Custodian | Regulatory Affairs Department |
MEDICAL CERTIFICATIONS
The compliance register is mapped to the following frameworks: EU MDR 2017/745, FDA 21 CFR Part 820 and 21 CFR Part 11, ISO 13485:2016, ISO 14971:2019, IEC 60601-1:2005+A1+A2, IEC 60601-1-2:2014+A1, IEC 62304, IEC 62366-1, UKCA UK MDR 2002 (as amended), Health Canada SOR/98-282, TGA Therapeutic Goods Regulations 1990, and ANVISA RDC 751/2022. Each certification body is assigned a re-certification horizon and a surveillance audit window.
CLINICAL ENVIRONMENT
The schedule is operationalized within the manufacturer’s quality management system and is reviewed at each management review cycle. Designated Regulatory Affairs personnel maintain the master register, while Clinical and Engineering functions receive impact notifications within five business days of a classified directive change. Annual compliance drills validate response readiness. The schedule is version-controlled, electronically signed, and archived for a minimum of ten years or the device lifetime, whichever is longer.
📥 Download Technical Specification
Click the button below to view or download the full official PDF datasheet.