
Recurso
Legal Liability Allocation Matrix in OEM Manufacturing – Official Clinical Overview & Technical Datasheet
EXECUTIVE SUMMARY
This whitepaper establishes the comprehensive Legal Liability Allocation Matrix for OEM manufacturing of medical aesthetic devices, delineating responsibilities across design, component sourcing, assembly, quality assurance, regulatory compliance, and post-market surveillance. The matrix serves as a contractual and operational reference for OEM partners, brand owners, and regulatory bodies, ensuring clarity in risk distribution and adherence to ISO 13485, FDA 21 CFR Part 820, and EU MDR 2017/745. By codifying liability boundaries, this framework mitigates disputes, enhances product safety, and streamlines market authorization processes.
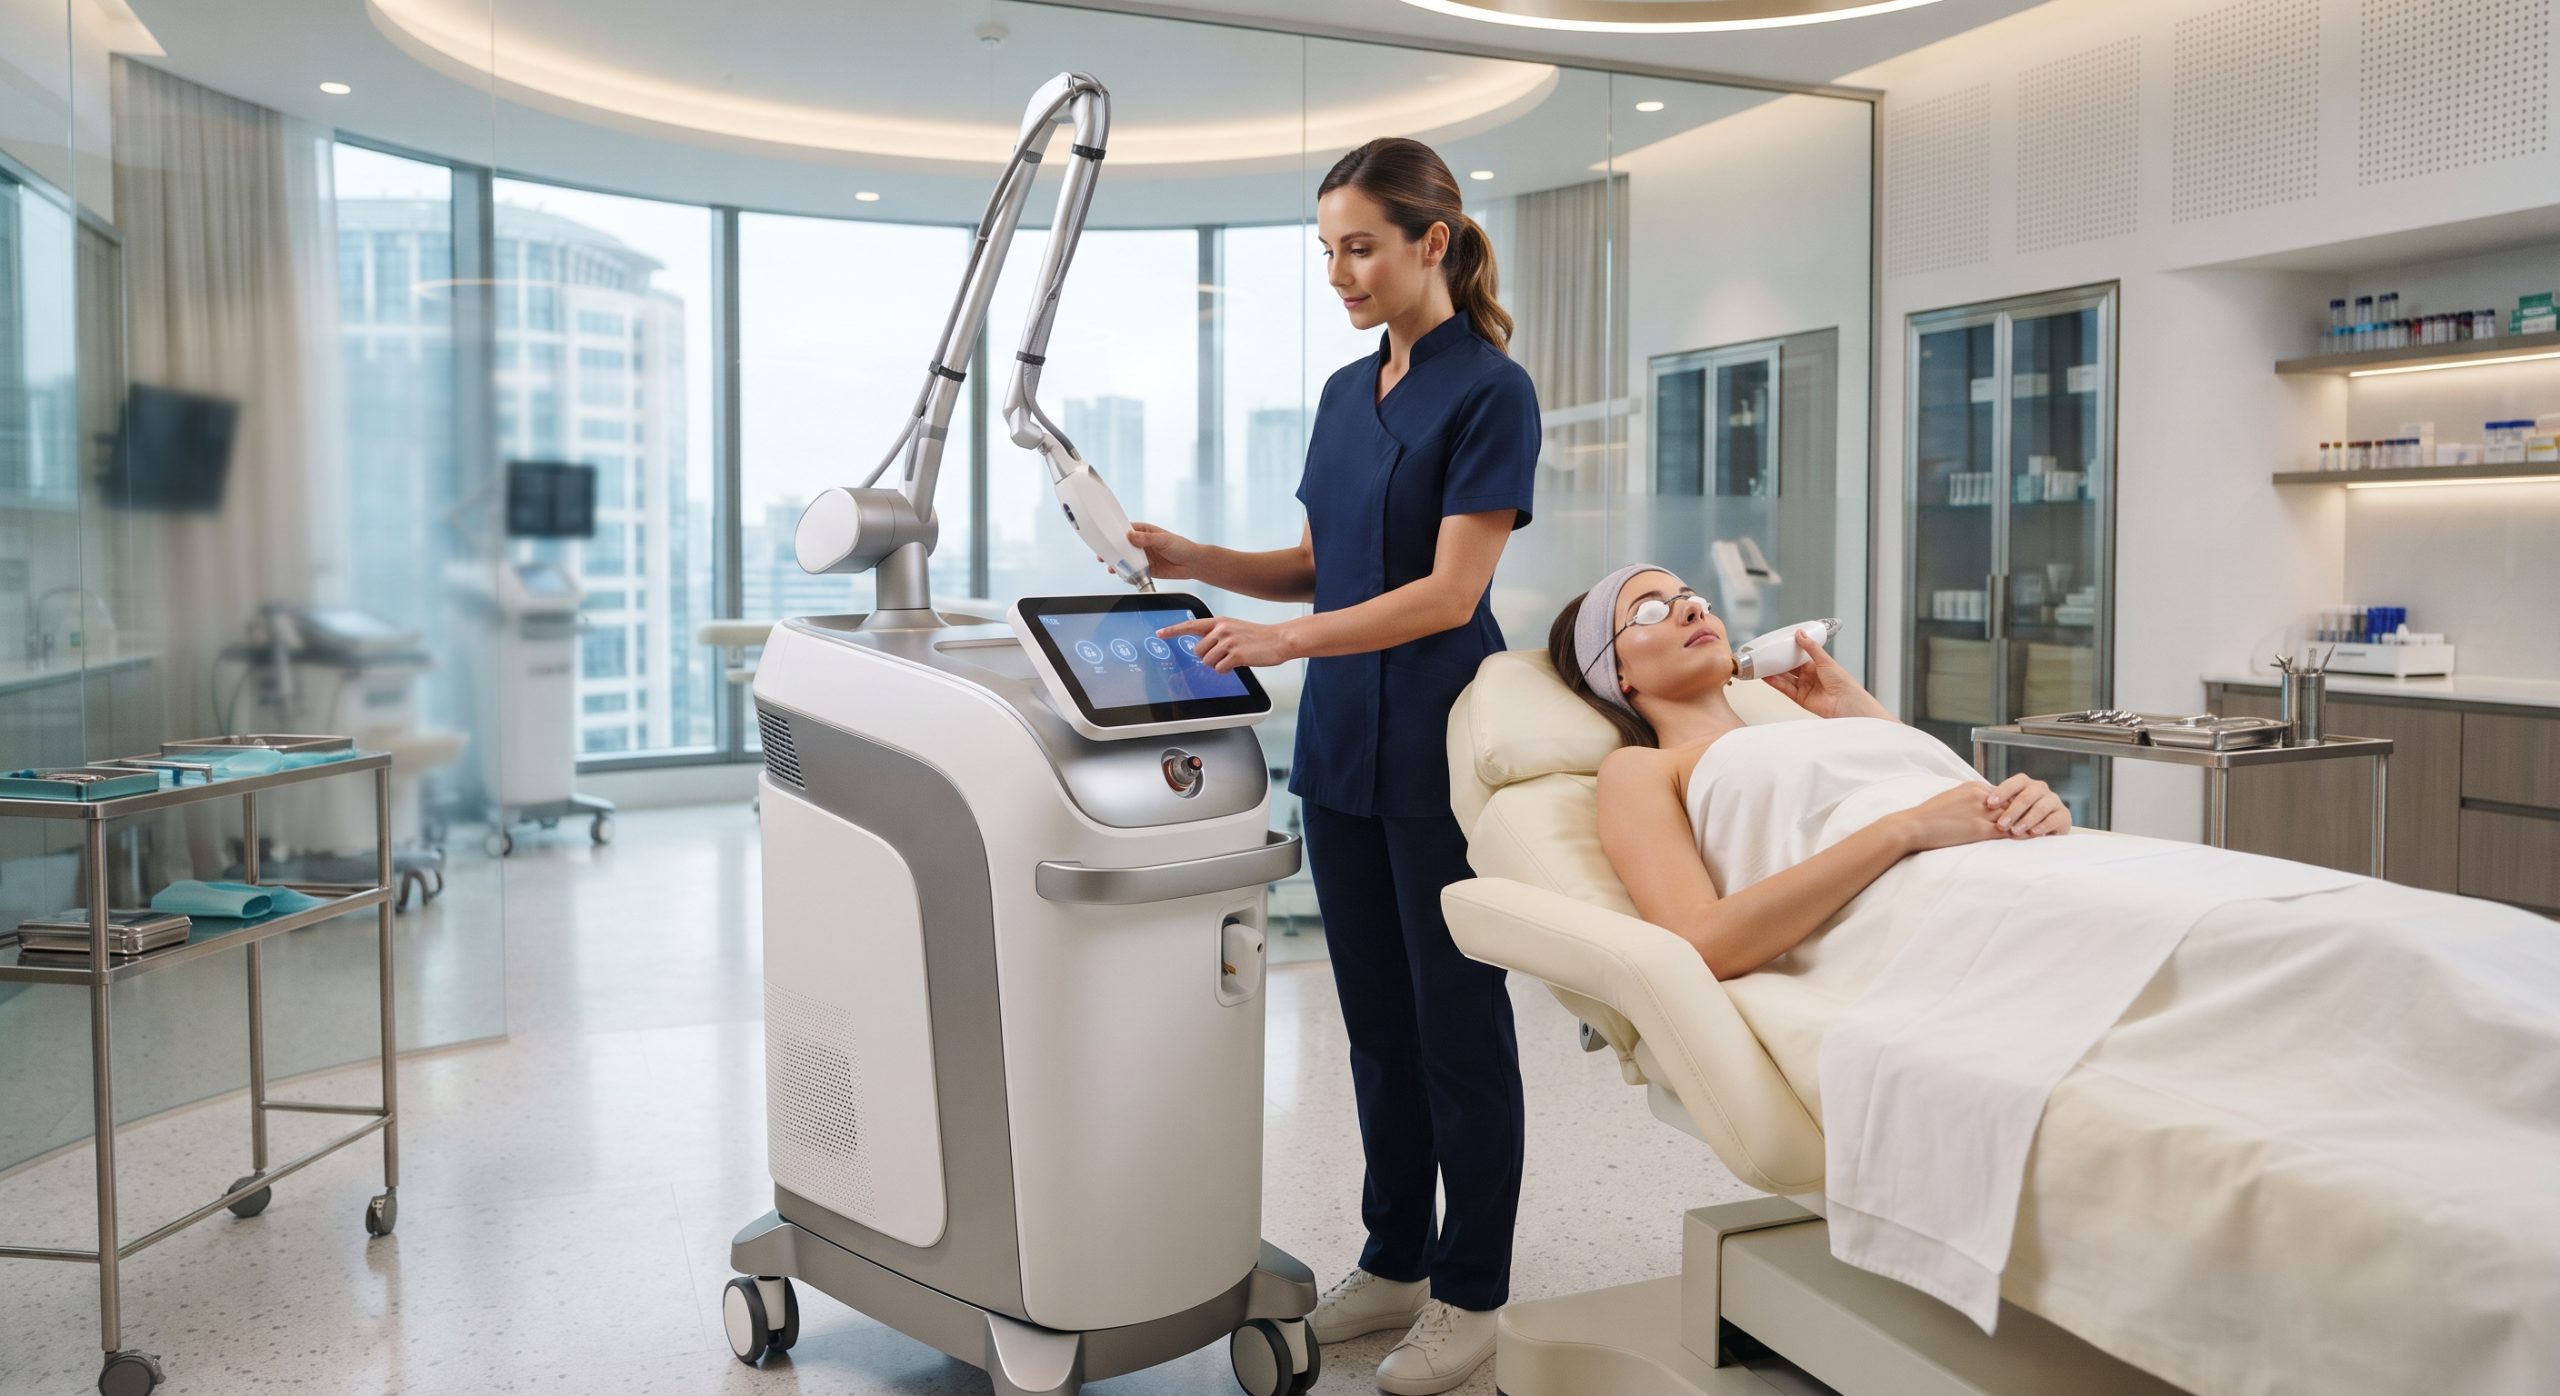
CLINICAL ARCHITECTURE & DESIGN
Liability allocation begins at the design phase. The OEM manufacturer assumes responsibility for design verification and validation, including electrical safety (IEC 60601-1), electromagnetic compatibility (IEC 60601-1-2), and software lifecycle management (IEC 62304). Brand owners retain liability for clinical claims, labeling, and indication-specific performance validation. The matrix requires joint sign-off on design history files (DHF) and risk management reports (ISO 14971), with explicit escalation protocols for design changes affecting safety or efficacy.
KEY INDICATIONS & CAPABILITIES
The liability matrix applies to all Class II and Class III aesthetic devices, including laser, RF, ultrasound, and cryolipolysis systems. Key capability areas addressed include optical source integrity, thermal safety thresholds, handpiece sterilization validation, and user interface fail-safes. Each capability is mapped to a responsible party: OEM for manufacturing defects, brand owner for misuse or off-label promotion, and shared liability for post-market clinical follow-up.
COMPLIANCE & STANDARDS
The matrix aligns with global regulatory frameworks: FDA Quality System Regulation (21 CFR 820), EU Medical Device Regulation (MDR 2017/745), ISO 13485:2016, ISO 14971:2019, and IEC 62366-1 usability engineering. Compliance verification requires documented traceability from component supplier to finished device, with liability assigned to the party responsible for the non-conformance. The OEM maintains audit readiness for notified body inspections; the brand owner ensures market-specific registrations and adverse event reporting.
TECHNICAL SPECIFICATIONS
[Text…]
| Liability Domain | OEM Responsibility | Brand Owner Responsibility | Shared Responsibility |
|---|---|---|---|
| Design & Development | Design verification, DHF, risk analysis | Clinical claims validation, labeling | Design change approval |
| Component Sourcing | Supplier qualification, incoming inspection | Approval of critical suppliers | Supplier audits |
| Manufacturing & Assembly | Process validation, workmanship | Packaging & labeling accuracy | Final release testing |
| Regulatory Compliance | ISO 13485, IEC 60601, CE technical file | Market registration, UDI, adverse event reporting | Post-market surveillance |
| Quality Assurance | Batch records, non-conformance handling | Complaint intake, CAPA initiation | Root cause analysis |
| Post-Market | Field corrective actions, recall execution | Customer notification, training updates | Trend reporting |
CLINICAL PROTOCOLS
Liability in clinical protocols is partitioned: the OEM provides validated treatment parameters and safety limits within the intended use; the brand owner is liable for clinical training, protocol customization, and patient selection criteria. Adverse event reporting follows a 24-hour notification chain from clinic to brand owner to OEM, with root cause analysis shared within 10 business days. Post-market surveillance data is jointly reviewed quarterly.
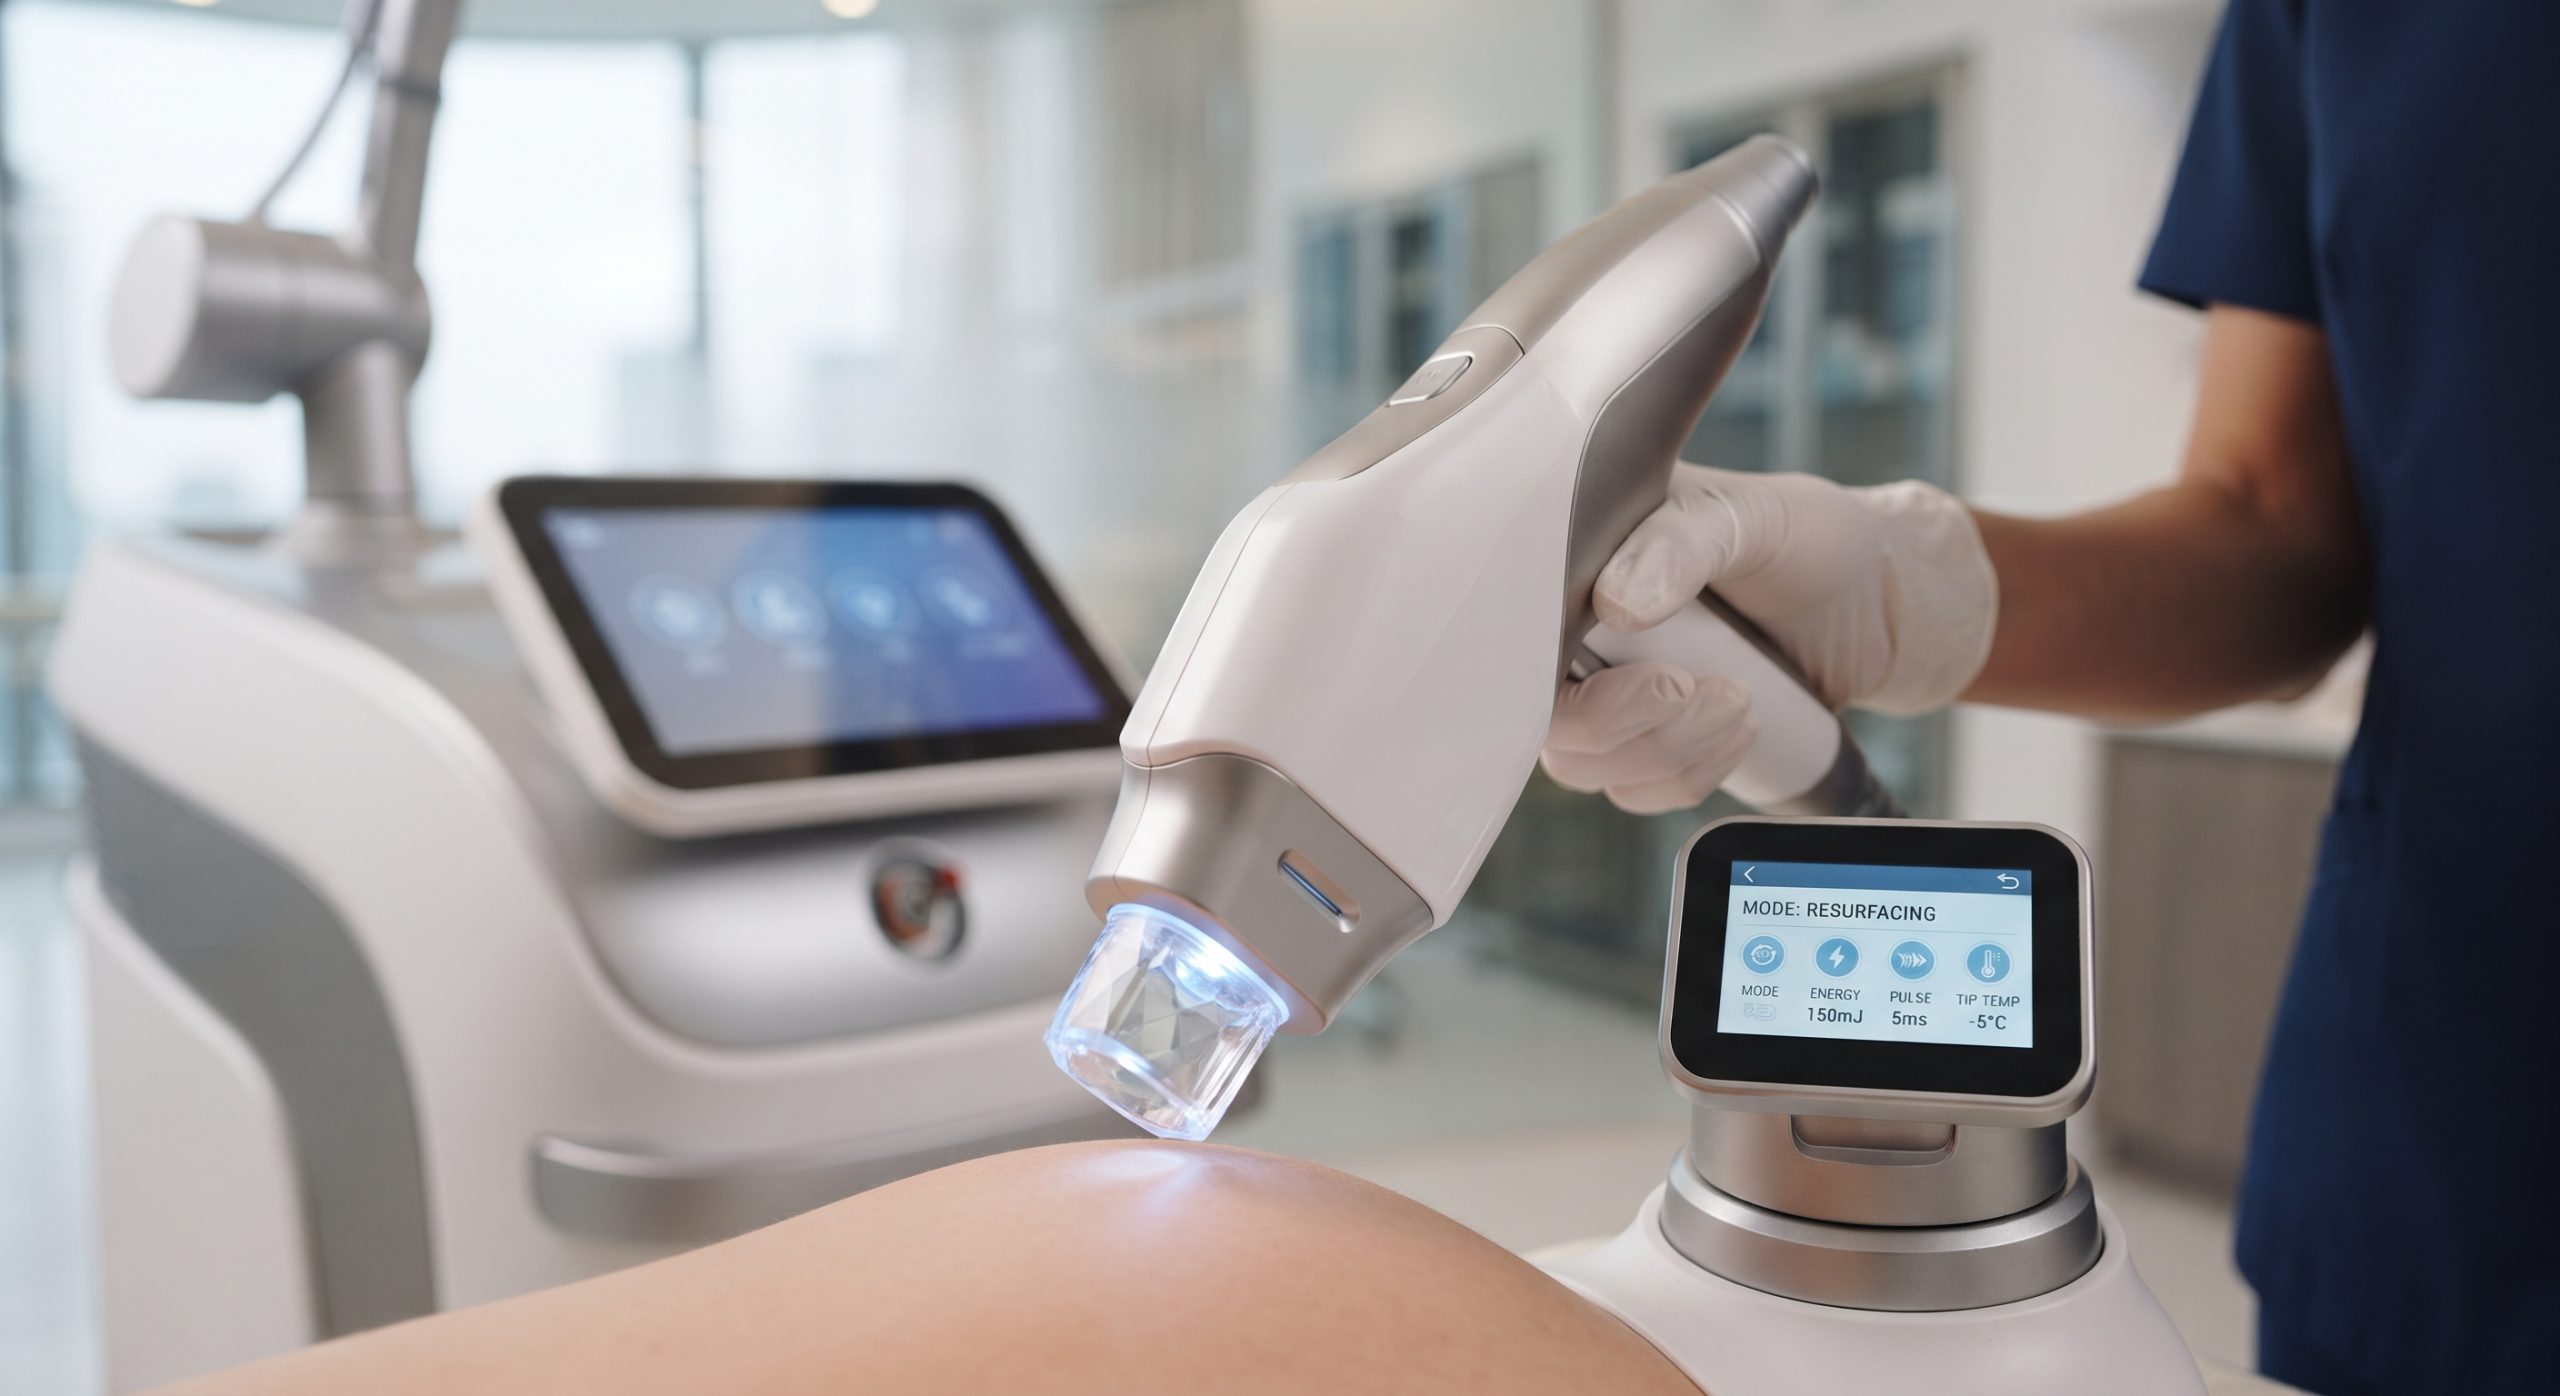
📥 Download Technical Specification
Click the button below to view or download the full official PDF datasheet.